nf-core/ncrnannotator












Introduction
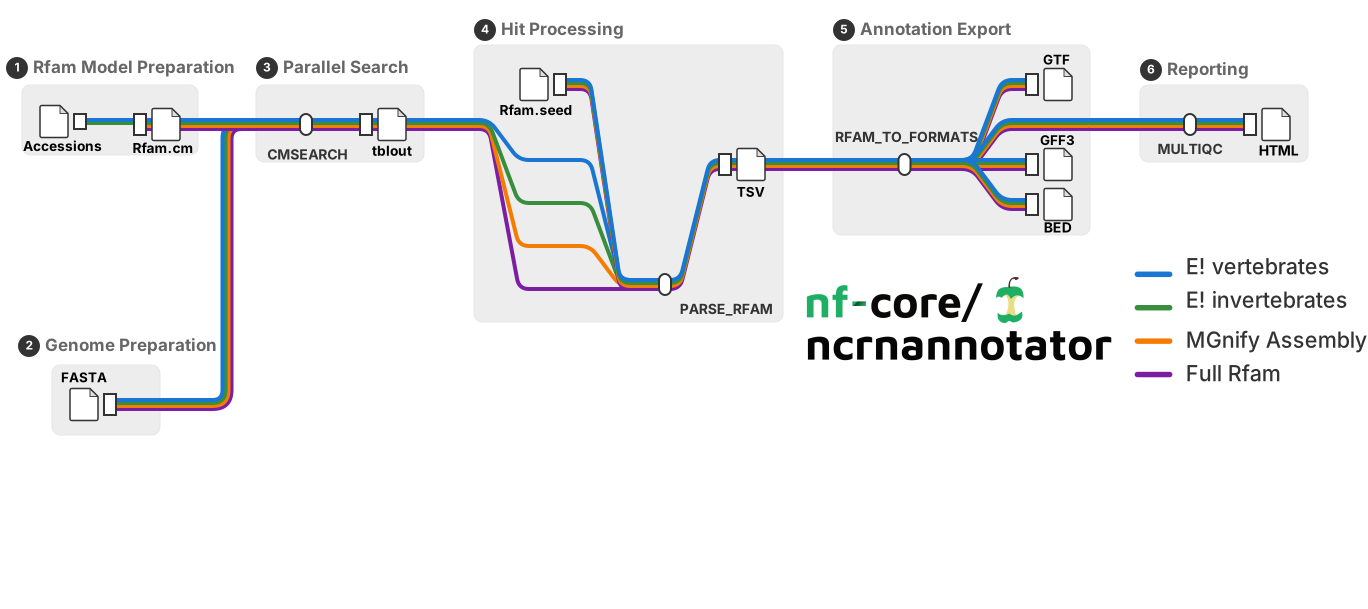
nf-core/ncrnannotator is a bioinformatics pipeline for genome-level non-coding RNA (ncRNA) annotation. It takes a genome assembly in FASTA format and annotates ncRNA loci using Rfam covariance models searched with Infernal (cmsearch). The pipeline is taxonomically aware, supporting vertebrate and invertebrate annotation modes, and produces output in GTF, GFF3, and BED formats.
- Filter Rfam covariance models to the target clade (
filter_rfam_cm) - Chunk genome into windows for parallel processing (
GENOME_CHUNK) - Search each chunk against filtered Rfam models (
Infernal cmsearch) - Parse hits, remove overlaps, apply GA score thresholds (
parse_rfam_results) - Export annotation in GTF, GFF3, and BED formats (
rfam_to_formats) - Present pipeline metrics (
MultiQC)
Usage
Note
If you are new to Nextflow and nf-core, please refer to this page on how to set-up Nextflow. Make sure to test your setup with -profile test before running the workflow on actual data.
ncrnannotator takes a genome assembly in FASTA format and Rfam database files as input. No samplesheet is required.
nextflow run nf-core/ncrnannotator \
  --fasta genome.fa \
  --mode ensembl-vertebrates \
  --rfam_cm Rfam.cm \
  --rfam_seed Rfam.seed \
  --outdir results \
  -profile docker
Warning
Please provide pipeline parameters via the CLI or Nextflow -params-file option. Custom config files including those provided by the -c Nextflow option can be used to provide any configuration except for parameters; see docs.
For more details and further functionality, please refer to the usage documentation and the parameter documentation. For more details and further functionality, please refer to the usage documentation and the parameter documentation.
Pipeline output
To see the results of an example test run with a full size dataset refer to the results tab on the nf-core website pipeline page. For more details about the output files and reports, please refer to the output documentation.
Credits
The first version of nf-core/ncrnannotator was written by Pedro Madrigal and Victoria Begley, members of the RNA Resources team at EMBL-EBI.
We thank the following people for their extensive assistance in the development of this pipeline:
Contributions and Support
If you would like to contribute to this pipeline, please see the contributing guidelines.
For further information or help, don't hesitate to get in touch on the Slack #ncrnannotator channel (you can join with this invite).
Citations
An extensive list of references for the tools used by the pipeline can be found in the CITATIONS.md file.
You can cite the nf-core publication as follows:
The nf-core framework for community-curated bioinformatics pipelines.
Philip Ewels, Alexander Peltzer, Sven Fillinger, Harshil Patel, Johannes Alneberg, Andreas Wilm, Maxime Ulysse Garcia, Paolo Di Tommaso & Sven Nahnsen.
Nat Biotechnol. 2020 Feb 13. doi: 10.1038/s41587-020-0439-x.
Generic options#
Less common options for the pipeline, typically set in a config file.
Display the help message.
Display version and exit.
Display the full detailed help message.
Custom logo file to supply to MultiQC. File name must also be set in the MultiQC config file
Email address for completion summary, only when pipeline fails.
More detail
An email address to send a summary email to when the pipeline is completed - ONLY sent if the pipeline does not exit successfully.
Custom config file to supply to M