nf-core/ribomsqc












Introduction
Use nf-core/ribomsqc to:
- Perform automated quality control of ribonucleoside analysis by mass spectrometry.
- Summarize and visualize QC metrics through integrated MultiQC reports.
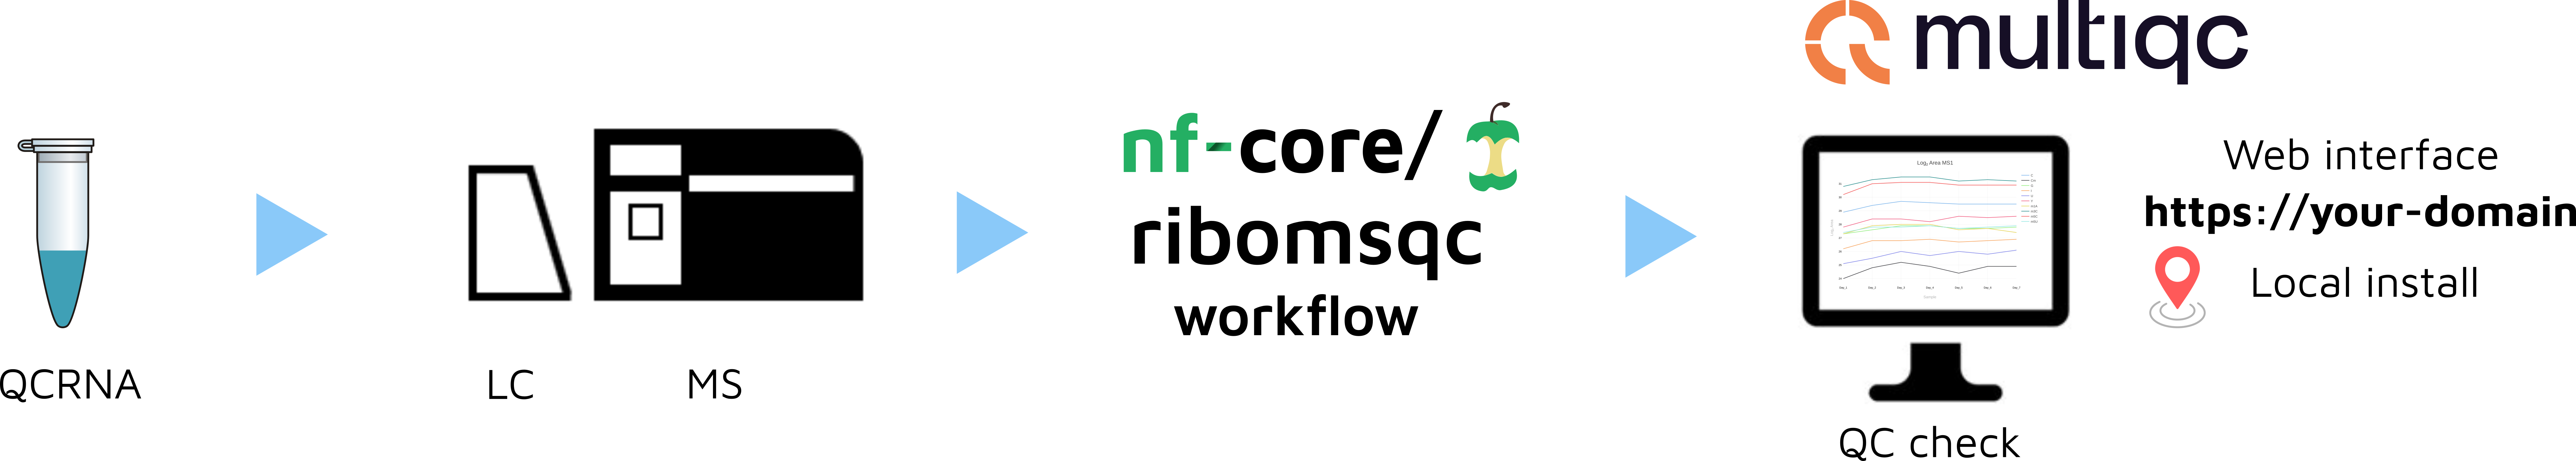 Figure 1: General workflow overview showing the pipeline steps from RAW file input through ThermoRawFileParser conversion, MSNBase XIC extraction, JSON merging, to final MultiQC report generation.
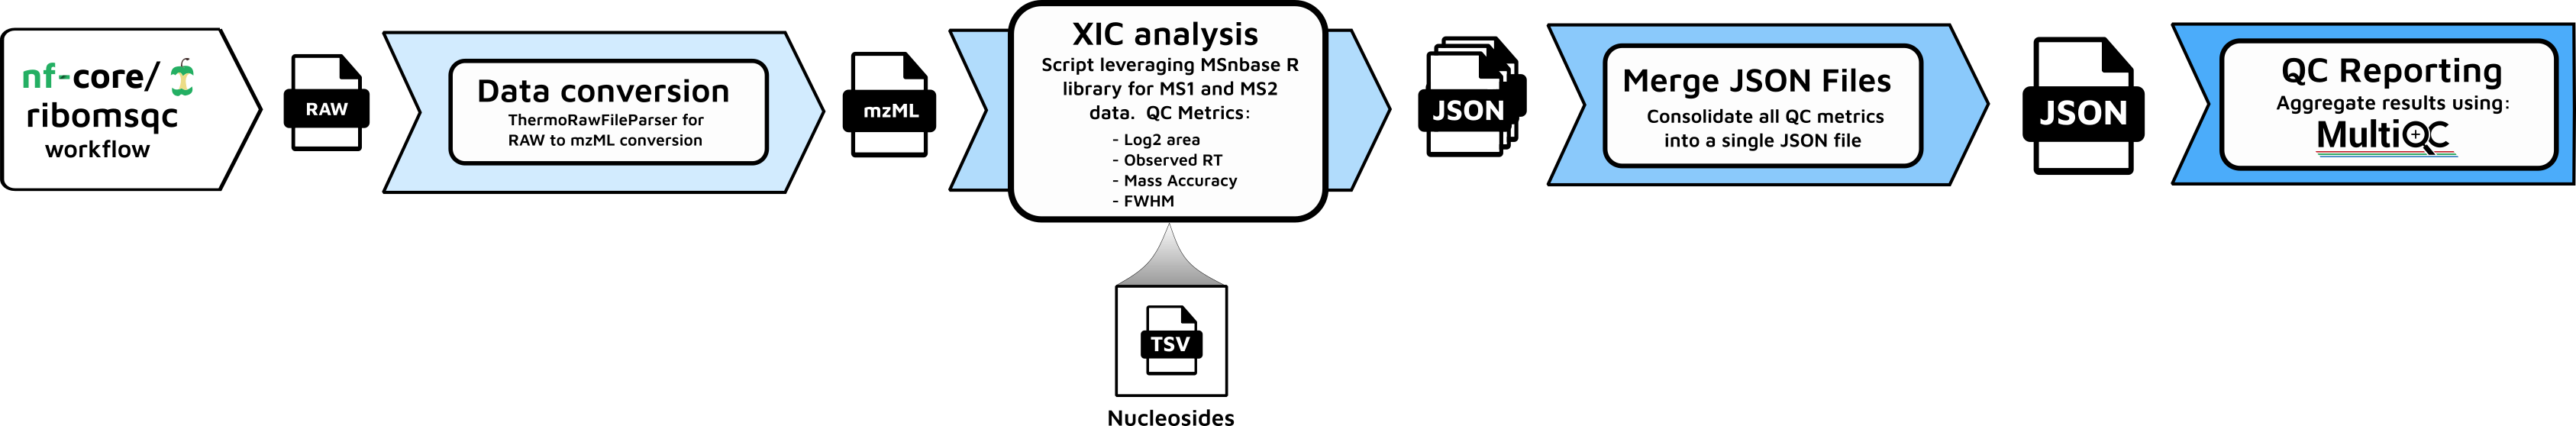 Figure 2: Detailed workflow diagram illustrating the data flow and process connections in the ribomsqc pipeline.
Usage
Note
If you are new to Nextflow and nf-core, please refer to this page on how to set-up Nextflow.
-
Prepare a samplesheet with your input data, for example:
id,raw_file
Day_5,path/to/Day_5.raw
For more information, see the usage docs on required samplesheet.csv columns.
- Prepare an analytes TSV file (e.g.
qcn1.tsv) with your compounds and theoretical retention times. The TSV must have exactly these columns and format:
short_name	long_name	mz_M0	mz_M1	mz_M2	ms2_mz	rt_teoretical
C	Cytidine 50 μg/mL	244.0928			112.0505	555
U	Uridine 25 μg/mL	245.0768			113.0346	1566
m3C	3-Methylcytidine methosulfate 100 μg/mL	258.1084			126.0662	508
m5C	5-Methylcytidine 100 μg/mL	258.1084			126.0662	655
Cm	2-O-Methylcytidine 20 μg/mL	258.1084			112.0505	883
m5U	5-Methyluridine 50 μg/mL	259.0925			127.0502	1866
I	Inosine 25 μg/mL	269.088			137.0458	1741
m1A	1-Methyladenosine 25 μg/mL	282.1197			150.0774	523
G	Guanosine 25 μg/mL	284.0989			152.0567	1726
m7G	7-Methylguanosine 25 μg/mL	298.1146			166.0723	554
Note
Replace only the values in the rt_teoretical column with your own retention times (in seconds) for each compound.
-
Run the pipeline:
nextflow run nf-core/ribomsqc \
  --input samplesheet.csv \
  --analytes_tsv qcn1.tsv \
  --analyte all \
  --rt_tolerance 120 \
  --mz_tolerance 7 \
  --ms_level 1 \
  --outdir results \
  -profile singularity
Warning
Please provide pipeline parameters via the CLI or Nextflow -params-file option.
For more information, see the usage docs and parameters.
Pipeline output
See results page for example output and output docs.
Credits
nf-core/ribomsqc was originally written by Roger Olivella.
Contributions and Support
For help, visit Slack #ribomsqc or see contributing guide.
Citations
See CITATIONS.md for tool references.
Ewels PA et al. (2020) The nf-core framework. Nat Biotechnol. doi:10.1038/s41587-020-0439-x
XIC extraction and plotting#
Parameters used for chromatographic peak extraction and visualisation
--analyte#
required
Short name of the analyte to be extracted, as defined in the 'short_name' column of the TSV file. Use a specific value such as 'm3C' to process one analyte, or use 'all' to process all analytes defined in the TSV file.
--ms_level#
required
MS level to extract chromatographic peaks from. Set to 1 for MS1 or 2 for MS2.
Enable test mode - pipeline continues even if analytes are not found in the data. Useful for CI testing with minimal datasets.
--analytes_tsv#
required
Path to a tab-separated values (TSV) file describing the analytes for chromatographic peak extraction. Must include columns: short_name, long_name, mz_M0 (required), and optionally mz_M1, mz_M2, ms2_mz, and rt_teoretical (required). Only mz_M0 and rt_teoretical are mandatory. mz_M1 and mz_M2 are reserved for future support of isotopic envelope integration.
--mz_tolerance#
required
Tolerance in parts-per-million (ppm) around the specified precursor m/z value (mz_M0) for XIC extraction. The window is defined as mz_M0 ± tolerance.
Whether to plot MS1 XICs
Whether to plot MS2 XICs
--rt_tolerance#
required
Time window (in seconds) around the theoretical retention time in which peaks will be searched. The window is defined as RT ± tolerance.